ANMAT: What Is Required in a Good Manufacturing Practices Inspection
21 Sep 2026 9 min read
ANMAT Disposition 4159/23 defines GMP requirements for pharmaceutical manufacturers in Argentina, covering quality systems, facilities, equipment, documentation, qualification, validation and quality control.
The Guide that Governs Medicine Manufacturing in Argentina, Annex by Annex
To understand how ANMAT guides Good Manufacturing Practices, one must follow the complete regulatory path: the authority defines national requirements, consolidates these requirements in Disposition 4159/23, incorporates international references such as WHO, PIC/S, ICH, and ISO standards, applies these criteria in the evaluation of facilities, equipment, documentation, production and quality control, and subsequently updates specific procedures, such as the certification of plants located abroad.
What is ANMAT?
ANMAT
is the national authority responsible for authorizing and inspecting companies that manufacture, package, import and export medicines in Argentina. In the manufacturing area, the document that concentrates its requirements is Disposition 4159/23, which approved the Guide to Good Manufacturing Practices for Manufacturers, Importers/Exporters of Medicines for Human Use. This guide defines how personnel, facilities, equipment, documentation, production and quality control should be, and is the text used as a reference to assess compliance during an inspection.
What international references guide ANMAT's GMP?
The ANMAT guide was not developed in isolation. According to the document itself, it incorporates international guidelines approved by the World Health Organization, reports from PIC/S (Pharmaceutical Inspection Co-operation Scheme), standards from ICH (International Council for Harmonisation) and ISO standards. The regulatory movement continued after its publication: in October 2025, ANMAT announced that, through
Disposition 7998/25
, the certification process for Good Manufacturing Practices for medicinal specialty plants located abroad was modernized, incorporating principles of Good Reliance Practices and establishing a 40-business-day deadline for evaluating initial documentation.
This content covers the structure of the guide and the points that appear most frequently in a laboratory's routine. You will find:
Which regulations were revoked by Disposition 4159/23 and why this matters
How the guide is organized into Part A, Part B and annexes
What ANMAT establishes about facilities, equipment and area qualification
What is the role of the Site Master File before ANMAT
What is reviewed in the annual product quality review
What happens when an inspection identifies recurring non-conformities
What ANMAT Disposition 4159/23 Revoked
Article 2 of the disposition revokes ANMAT Dispositions No. 2819/04, 3602/18, 3827/18 and 1281/19. This means that the set of general Good Manufacturing Practices guidelines in force since 2004 and their subsequent updates was replaced by a single text. Article 3 established that the disposition would enter into force after 30 business days.
For a laboratory, the practical effect is simple, albeit relevant: any internal procedure, checklist or reference document that cited the revoked regulations now presents an outdated regulatory reference, even when its technical content remains valid.
How the ANMAT Good Manufacturing Practices Guide Is Structured
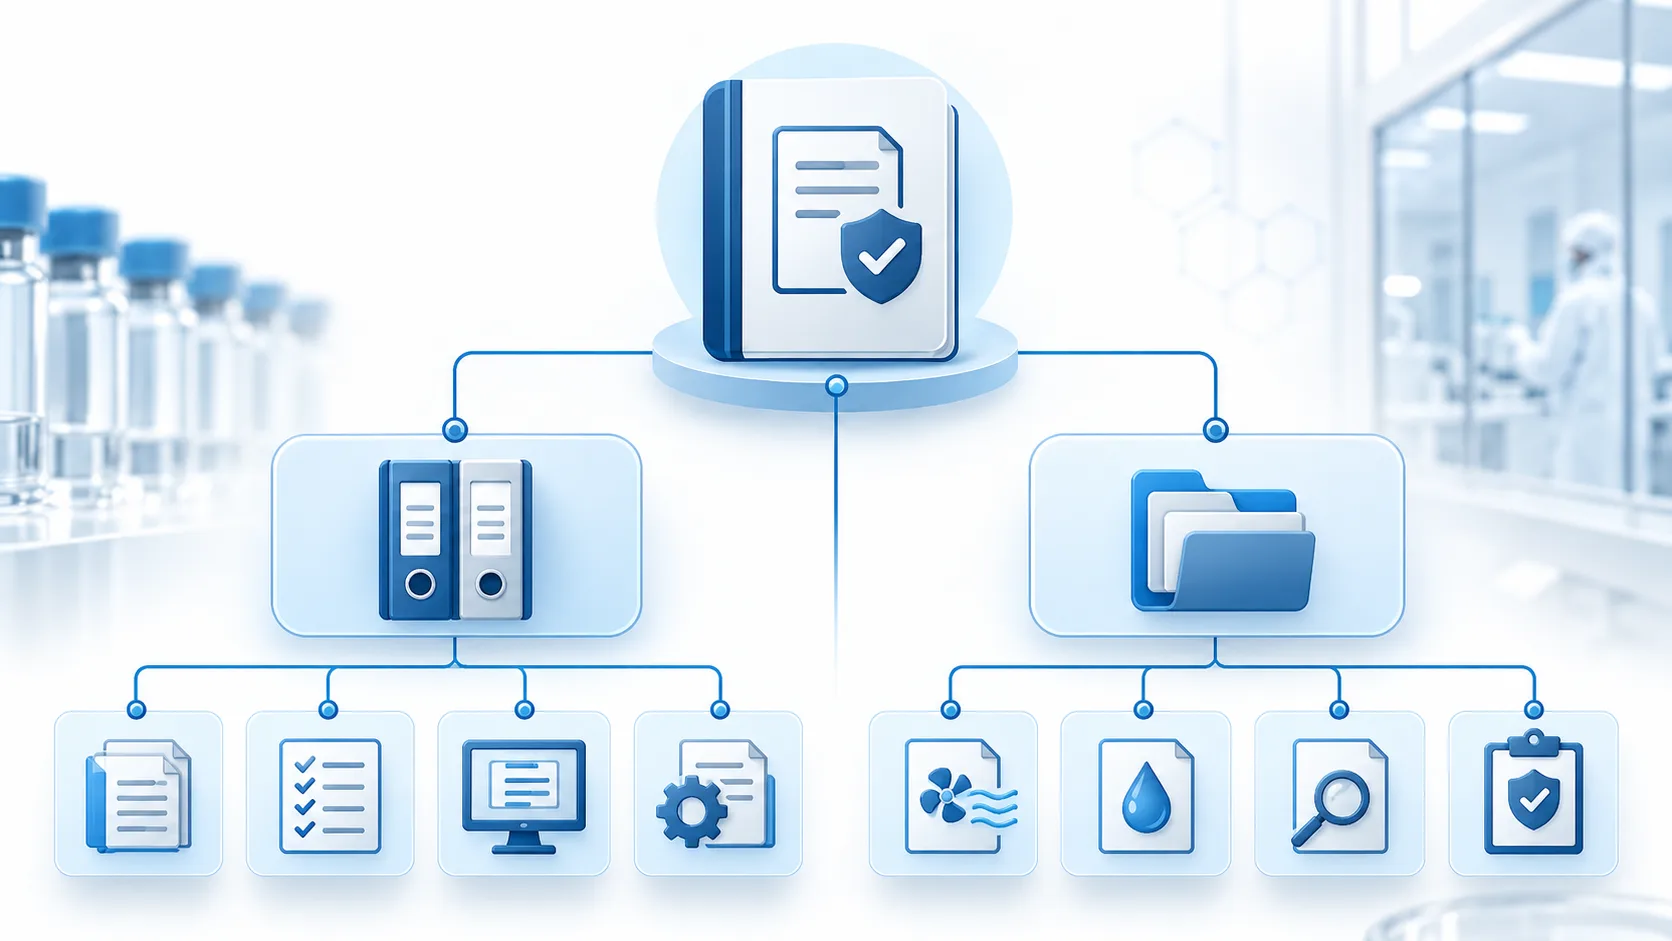
The guide is divided into two parts and nineteen annexes. Part A brings together the basic requirements for manufacturing, importing and exporting medicines, organized into ten chapters: Pharmaceutical Quality System, Personnel, Facilities and Equipment, Documentation, Production, Quality Control, Outsourced Activities, Complaints, Quality Defects and Product Recalls, Self-Inspection and Glossary. Part B addresses the basic requirements for Active Pharmaceutical Ingredients (APIs) used as starting materials.
The annexes address topics that were previously dispersed or not directly covered.
Annex 1:
manufacture of sterile medicines
Annex 2:
sampling of starting and packaging materials
Annex 3:
reference and retention samples
Annex 5:
qualification and validation
Annex 6:
computerized systems
Annex 7:
water for pharmaceutical use
Annex 8:
quality risk management
Annex 9:
air handling systems for non-sterile product areas
The list also includes the use of ionizing radiation, parametric release, manufacture of liquids and semi-solids, pressurized aerosols for inhalation, medicinal gases, herbal medicines, biological medicinal products, medicines derived from human blood or plasma and radiopharmaceuticals. This structure explains why ANMAT may apply different inspection criteria to two plants that manufacture different pharmaceutical forms.
Facilities, Equipment and Area Qualification According to ANMAT
Requirements for Facilities and Equipment
Chapter 3 of the guide establishes that facilities and equipment must be located, designed, constructed, adapted and maintained in a manner appropriate to the operations performed, minimizing the risk of errors and allowing effective cleaning and maintenance.
Area Qualification and ISO 14644
Item 3.12 establishes that production areas must have effective ventilation, with air control installations appropriate to the products and operations performed; that, for the qualification of areas, the guidelines of ISO 14644 standards or higher must be followed; and that monitoring of conditions must be performed using calibrated instruments.
The reference to ISO 14644 appears again in Chapter 5, dedicated to production. Among the technical measures aimed at preventing cross-contamination, item 5.21 mentions independent production areas, with separate process equipment and HVAC systems, and determines that the qualification of areas, according to the products handled or manufactured, must follow the guidelines of ISO 14644 or another document that describes similar or superior methodology and requirements.
Calibration and Verification of Equipment
Item 3.41, in turn, establishes that measurement, weighing, recording and control equipment must be calibrated and verified at defined intervals, using appropriate methods, and that adequate records of these verifications must be maintained.
Requirements for Quality Control Laboratories
In the control laboratory, the guide also establishes an external reference. Chapter 6 determines that laboratories intended for Quality Control must comply with the guidelines of ISO 17025.
The Site Master File and the Documentary Relationship with ANMAT
What is the Site Master File (SMF)?
Chapter 4 of the guide, when listing the required GMP documentation, defines the Site Master File, Archivo Maestro de Sitio (AMS), as the company document that describes activities related to Good Manufacturing Practices and refers to the specifications set forth in ANMAT Disposition No. 7066/2013 or its amendments.
That disposition established that companies authorized by ANMAT to manufacture or package medicines, in their own or third-party plants, as well as to import or export medicines or active pharmaceutical ingredients, must submit the Site Master File annually. According to the recitals of the regulation, the SMF must provide specific and clear information on the management of quality policies and production and quality control activities, in addition to meeting ANMAT's need to have updated technical, administrative and legal information on each authorized company in order to program its oversight more effectively.
How is the Site Master File submitted to ANMAT?
The process was also modernized. ANMAT announced the implementation of the submission of the Site Master File according to Disposition 7066/2013 through the
TAD platform
, before the National Institute of Medicines (INAME).
The Annual Product Quality Review
Item 1.10 of the guide requires periodic or continuous quality reviews of all authorized medicines, including those intended exclusively for export, with the objective of verifying process consistency, evaluating the adequacy of current specifications, identifying trends and detecting opportunities for improvement. These reviews must be performed and documented, generally annually.
What should be evaluated in the quality review?
The list of minimum contents of this review is extensive and includes critical in-process controls, non-conforming batches and their respective investigations, significant deviations and the effectiveness of corrective and preventive actions, changes in processes or analytical methods, results of the ongoing stability program, and returns, complaints and market recalls.
Among these points, one connects directly to the laboratory's infrastructure: subsection xi requires the review of the qualification status of major equipment and services, expressly mentioning air handling systems (HVAC), water and compressed gases.
What Happens When ANMAT Identifies Non-Conformities
A Recent Case: ANMAT Disposition 2990/2026
The consequences of a non-conformity can become public. In May 2026, the
Official Gazette
published ANMAT Disposition 2990/2026, which details that, during a follow-up inspection, deficiencies related to compliance with Good Manufacturing and Control Practices according to ANMAT Dispositions No. 4159/23 and 7298/19 were identified, most of which were recurrences of non-conformities identified in previous inspections.
The text enumerates the specific items of Part B considered non-compliant and describes findings such as the absence of a robust, defined and documented Pharmaceutical Quality System, lack of adequate resources for its implementation or maintenance, procedures that did not cover all operations, and personnel without adequate training. Based on these points, INAME suggested the preventive inhibition of the company's production activities and import and export operations.
The most relevant point of this document is not the specific case, but the method used: ANMAT cites exactly the item of the guide that was breached. This transforms Disposition 4159/23 into a document that must be known not only conceptually, but also by its specific items.
Frequently Asked Questions about ANMAT and Good Manufacturing Practices
What is ANMAT?
It is the Administración Nacional de Medicamentos, Alimentos y Tecnología Médica, the body responsible for authorizing and inspecting companies that manufacture, import and export medicines in Argentina.
What is the current Good Manufacturing Practices disposition from ANMAT?
Disposition 4159/23, which approved the Guide to Good Manufacturing Practices for Manufacturers, Importers/Exporters of Medicines for Human Use and revoked Dispositions 2819/04, 3602/18, 3827/18 and 1281/19.
Which international standard does ANMAT cite for area qualification?
ISO 14644, mentioned in both item 3.12 and item 5.21 of the guide. The disposition also accepts other documents that describe similar or superior methodology and requirements.
Does the guide require calibrated equipment?
Yes. Item 3.12 establishes that monitoring of conditions must be performed with calibrated instruments, while item 3.41 requires that measurement, weighing, recording and control equipment be calibrated and verified at defined intervals, with maintenance of the respective records.
How often must the Site Master File be submitted to ANMAT?
Annually, according to Disposition 7066/2013. The submission was implemented through the TAD platform before INAME.
Which standard applies to quality control laboratories?
Chapter 6 of the guide establishes that laboratories intended for Quality Control must comply with the guidelines of ISO 17025.
Which annex deals with water for pharmaceutical use?
Annex 7. Item 5.65 of the guide refers to this annex by establishing that water used in the manufacture of pharmaceutical products must have quality appropriate to the intended use.
What happens when an inspection identifies recurring non-conformities?
The measures adopted may be published in the Official Gazette. In the most serious cases, INAME may suggest the preventive inhibition of the company's production activities and import and export operations.
Related Readings
ISO 14644: classification of air cleanliness in cleanrooms
Qualification and validation of critical systems
Water for pharmaceutical use: types of water and controls